Supplier qualification that controls what purchasing can buy
For QA and procurement heads · Compare approaches

Supplier qualification that controls what purchasing can buy: Connecting supplier qualification status directly to purchasing controls ensures procurement orders only flow to approved active-ingredient and excipient sources.
Quick answer: supplier qualification pharma
Connect suppliers, manufacturing sites and materials to approved purchasing scope. Use a worked example, exception checks and practical pilot actions. Learn why an approved supplier name is insufficient when materials can come from different manufacturing sites.
TL;DR — what you’ll learn
- Learn why an approved supplier name is insufficient when materials can come from different manufacturing sites.
- Build an approval record that connects supplier, manufacturer, site, material and specification.
- See how purchasing can check that record without taking over the authorised quality team's qualification decision.
- Use the worked example and pilot checks to identify where your current process can permit an unsupported purchase.
A buyer selects a familiar supplier. The purchase order passes the normal commercial approvals, and the material arrives on time. Receiving then discovers that the manufacturer has supplied it from a different site. The vendor is approved in the enterprise resource planning (ERP) system, but the site-material combination has never been assessed for the intended use.
The useful control is an approval with a defined scope. Purchasing needs to know which source is acceptable for which material and use, under which conditions, at the time the order is placed. A single green flag against a vendor cannot express that decision reliably.
For Indian pharmaceutical operations, start from the applicable requirements and approved quality procedures. Revised Schedule M includes supplier evaluation and approved-source controls. Its requirements differ across relevant parts, so the assessment must identify the applicable provision rather than applying one generic checklist everywhere. Part I section 10.8 assigns supplier approval to the person responsible for quality control together with relevant departments; reflect that applicable authority in the workflow. See Schedule M, including Part I section 10.8 and API materials controls.
Define the object being approved
Commercial supplier, original manufacturer and manufacturing site may be different entities. A distributor can supply materials from several manufacturers. A manufacturer can operate multiple sites. The same site can make several grades of a material, with different specifications and intended uses.
Represent those relationships explicitly. The commercial supplier remains the entity against which purchasing and payment occur. The manufacturing source identifies where the material originates. The approval connects the source to the material or approved material family and its permitted use.
This is an engineering model to support the organisation's quality decision. The responsible quality function should determine the necessary granularity from the material, process and regulatory context. Do not create thousands of unnecessary approval combinations simply because the software can store them. Equally, do not compress materially different sources into one code merely to make data entry easier.
For API operations, ICH Q7 materials-management guidance, section 7, is a useful reference for supplier evaluation and source controls. It is API guidance; its scope should remain visible when a team uses it alongside Indian requirements.
Work through the approval combinations
Consider an illustrative business buying three materials from one supplier. The supplier uses two manufacturing sites. This produces six possible site-material combinations, but the quality team has approved only two.
Scroll horizontally to see all columns.
| Manufacturing site | Material | Current approved scope | Purchasing response |
|---|---|---|---|
| Site A | Material X | Approved for the stated specification and use | Permit within that scope |
| Site A | Material Y | No approval recorded | Route to qualification owner |
| Site A | Material Z | No approval recorded | Route to qualification owner |
| Site B | Material X | No approval recorded | Route to qualification owner |
| Site B | Material Y | Approved for the stated specification and use | Permit within that scope |
| Site B | Material Z | No approval recorded | Route to qualification owner |
The supplier's commercial status does not resolve the four missing combinations. Neither does a purchase order below a financial approval threshold. The system should explain the specific gap: “Site B is not approved for Material X under this specification,” rather than displaying an unhelpful “vendor blocked” message.
Suppose Site A later loses approval for Material X. That change should trigger assessment of relevant open orders and incoming shipments. It should not silently erase the earlier approval or rewrite the history of previous purchases. The organisation needs to reconstruct what was known and authorised at each decision point.
Give the qualification record a usable lifecycle
Start with the request to qualify a source. Record the business need, proposed material, source identity, specification and responsible quality owner. Procurement supplies commercial and logistical information; quality determines the evidence and assessment needed for the proposed use.
A practical workflow distinguishes at least these business states:
- Requested: the source is proposed and the reason is recorded.
- Assessment in progress: evidence and identified gaps are being reviewed.
- Approved within scope: the authorised decision and its conditions are recorded.
- Restricted or suspended: a defined limitation prevents some or all use.
- Closed or superseded: the record is retained but no longer governs new activity.
These are suggested system states, not a mandatory regulatory vocabulary. Match them to the approved procedure. If conditional approval exists in that procedure, record the exact condition and permitted activity. A label called “conditional” is unsafe as a business control when users cannot tell whether it permits ordering, receipt, testing or production use.
Retain the evidence reviewed, the decision maker, approval date, effective date, scope and any review trigger. A scheduled review date and an approval expiry date are different concepts. Do not automatically suspend a valid source merely because an informational reminder is late unless the approved procedure requires that outcome.
Collect the fields needed for a purchase decision
The following record is a starting point for a design workshop:
Scroll horizontally to see all columns.
| Field | Question it answers |
|---|---|
| Commercial supplier and original manufacturer | Who sells the material, and who makes it? |
| Manufacturing site identifier | Which physical source does the assessment cover? |
| Material code, grade and specification version | What exactly may be purchased? |
| Intended product, process or use scope | Where is the approval applicable? |
| Status, effective dates and restrictions | Is the proposed activity permitted now? |
| Evidence references and quality owner | Where can the decision be checked? |
| Change and review history | What changed, and what needs reassessment? |
Use controlled identifiers where records must join across systems. Keep supplier aliases available for searching, but do not let a similar company name establish that two sources are identical. Where legacy data is incomplete, show the missing field and its owner. Replacing an unknown site with the supplier's head-office address creates false certainty.
Connect the decision to purchasing and receiving
A useful purchasing check runs against the actual order line. It compares the requested material, specification and source with the effective approval scope. If the source is not yet known, the order should follow a defined exception path rather than assuming the vendor's default site.
Repeat the relevant check at receiving. An order can be correct while the delivered source differs. Receiving should capture the source information available on the documents and labels and route discrepancies to the responsible team. It should not edit source identity simply to make the receipt match the order.
Decide which system owns approval. Often the quality workflow holds the authoritative decision while ERP consumes a controlled status or approval reference. The integration needs a version or effective timestamp, an acknowledgement and a reconciliation queue. A failed update must be visible; otherwise purchasing may continue using yesterday's status.
Keep commercial approval separate. Procurement can approve price and delivery terms while the designated quality owner retains the source qualification decision. Both decisions may be necessary, but neither substitutes for the other. The purchase-approval guide develops that distinction.
Test the exceptions before expanding the scope
Pilot one material family with enough variation to exercise the real process. Include an approved manufacturer supplied through a distributor, a second manufacturing site, a changed specification and an order created before a suspension.
Ask the team to demonstrate these outcomes:
- An approved combination is accepted for the permitted activity.
- An unsupported combination is stopped or routed according to procedure, with a specific reason.
- A later source restriction identifies relevant open transactions for assessment.
- A stale integration message cannot overwrite a newer approval decision.
- Receiving can report a source mismatch without changing the original order history.
- An authorised reviewer can reconstruct the evidence and approval effective at purchase time.
Measure the number of unresolved source combinations, time spent obtaining missing evidence and purchases requiring rework. Do not evaluate the project solely by the percentage of automatically approved orders. A system that routes genuine uncertainty correctly can be more useful than one that reports a high pass rate by ignoring missing data.
AI can help classify documents, extract proposed identifiers or identify a possible inconsistency. The extracted information should remain linked to its source and enter a reviewed workflow. A model-generated supplier score is not a qualification decision.
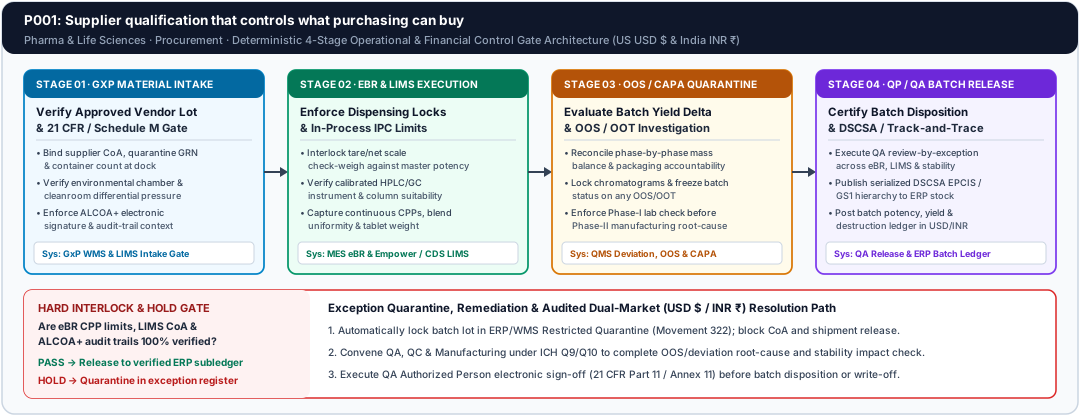
Deterministic State-Machine Architecture and 21 CFR Part 11 Interlocks
To resolve the core operational breakdown in supplier qualification pharma—specifically where a supplier is approved at company level but its new manufacturing site is not covered.—QA and procurement heads leaders must replace passive post-hoc reviews with an active state-machine contract across ERP Materials Management (SAP MM / WM), Warehouse Management System (WMS), LIMS Incoming QC Queue, and L1–L5 Serialization / EPCIS Repository. In a hybrid or paper-dependent plant, operators and analysts can inadvertently advance a batch or analytical sequence from Separate supplier from site and material to Define qualification evidence even when a prerequisite calibration, vendor status, or environmental check has expired.
Under a deterministic GxP stage-gate architecture, every transaction in approved supplier list is cryptographically bound to Vendor Site ID, Material Code (MATNR), Goods Receipt Batch (CHARG), CoA Checksum, and GS1 SSCC / SGTIN-96 identifier. If the automated validation rule at Define qualification evidence detects an out-of-tolerance parameter, a missing second-person verification signature, or an unlinked source artifact, the middleware immediately transitions the lot into a hard RESTRICTED-QA-HOLD state across ERP and MES/LIMS, routing the exception into Control approval scope and expiry before any downstream processing or batch certification can occur.
Scroll horizontally to see all columns.
| Stage-Gate Transition in Supplier qualification that controls what purchasing can buy | Authoritative System Boundary | Mandatory ALCOA+ Payload & Identity Key | Automated Quarantine & Escalation Rule |
|---|---|---|---|
| Gate 01: Separate supplier from site and material | ERP Materials Management (SAP MM / WM) | Active Vendor Site ID and authenticated operator/analyst ID | Reject transaction initiation if master data, equipment, or vendor status is not RELEASED |
| Gate 02: Define qualification evidence | Warehouse Management System (WMS) | Contemporaneous UTC timestamp, raw instrument telemetry, and specification token | Block progression and trigger immediate alert if value breaches warning/action limits |
| Gate 03: Control approval scope and expiry | Enterprise eQMS (Deviation / OOS / CAPA) | Linked parent-child batch genealogy array, ICH Q9 risk score, and root-cause code | Lock ERP inspection lot (QA11) and prevent bypass until investigation is verified |
| Gate 04: Worked example and usable record or workflow | QA Release & Part 11 Archive Vault | Dual electronic signature (21 CFR Part 11 / Annex 11) and SHA-256 audit packet | Release batch to UNRESTRICTED stock only when 100% of gate predicates evaluate to TRUE |
Tri-Jurisdictional Inspection Readiness: US FDA, EU Annex 11, and Revised Schedule M
Pharmaceutical manufacturers operating across domestic and regulated export markets must engineer supplier material site approval so a single electronic workflow satisfies US FDA, European EMA/PIC/S, and Indian CDSCO Revised Schedule M inspectors simultaneously. While regional guidance documents use different clause numbering, all three regulatory authorities converge on the same non-negotiable requirement: contemporaneous, attributable, and tamper-evident control over supplier qualification that controls what purchasing can buy.
- US FDA cGMP & 21 CFR Part 11 Expectations: Under 21 CFR § 211.80–211.94 (Control of Components, Containers, and Closures), FDA Drug Supply Chain Security Act (DSCSA) EPCIS 1.2 unit-level serialization, and ICH Q7 Section 7 (Materials Management), FDA investigators routinely request raw electronic sequences, dynamic audit trails, and exception logs for Supplier qualification that controls what purchasing can buy rather than relying on printed summary PDFs. Any discrepancy between physical shop-floor/laboratory execution and electronic timestamps—or any uninvestigated abort/override—is cited under 21 CFR § 211.192, § 211.194, or Part 11.10(e).
- EU GMP, PIC/S PI 041-1 & Qualified Person (QP) Certification: Under EU GMP Chapter 5 (Production & Starting Materials), EU Annex 16 (QP Batch Certification & Supply Chain Traceability), and EU Falsified Medicines Directive (FMD), European inspectors and Qualified Persons require structured Review-by-Exception (RbE) evidence demonstrating that computerized systems enforce Annex 11 Clause 12.4 audit-trail review and that all supply-chain, manufacturing, and analytical controls align with the registered Marketing Authorization (MA) dossier.
- India CDSCO Revised Schedule M (G.S.R. 922(E)) & WHO GMP: Under CDSCO Revised Schedule M (G.S.R. 922(E) Part I Section 10–11 Materials & Warehousing), WHO TRS 1025 Good Storage and Distribution Practices (GSDP), and DGFT/iVEDA barcoding & cold-chain rules, Indian formulation and API plants must operate a digital Pharmaceutical Quality System (PQS) that links every deviation, OOS/OOT, vendor change, and stability trend directly to batch disposition and Annual Product Quality Review (APQR) reporting.
Scroll horizontally to see all columns.
| Regulatory Authority | Governing Statutory & Guidance Clauses | Primary Audit Inspection Vector for Supplier qualification that controls what purchasing can buy | Built-In System Defense & Evidence Packet |
|---|---|---|---|
| United States (`US FDA`) | 21 CFR § 211.80–211.94 (Control of Components, Containers, and Closures), FDA Drug Supply Chain Security Act (DSCSA) EPCIS 1.2 unit-level serialization, and ICH Q7 Section 7 (Materials Management) | Contemporaneous raw data capture, Part 11 audit trails, and OOS/deviation rigor | Append-only audit log capturing Actor_ID, UTC_Time, Old_Val, New_Val, Reason |
| European Union (`EMA / PIC/S`) | EU GMP Chapter 5 (Production & Starting Materials), EU Annex 16 (QP Batch Certification & Supply Chain Traceability), and EU Falsified Medicines Directive (FMD) | Annex 11 computerized validation, Annex 15/16 QP certification, and ICH Q9 risk gates | Automated RbE exception summary bound to batch release certificate |
| India & WHO (`CDSCO Schedule M`) | CDSCO Revised Schedule M (G.S.R. 922(E) Part I Section 10–11 Materials & Warehousing), WHO TRS 1025 Good Storage and Distribution Practices (GSDP), and DGFT/iVEDA barcoding & cold-chain rules | End-to-end batch traceability, vendor/material locks, and CAPA effectiveness | Single-click genealogy and financial/quality reconciliation across all plant sites |
Right-First-Time (RFT) Yield and Dual-Market Working-Capital Impact
For QA and procurement heads executives and plant heads, enforcing automated stage gates in supplier qualification pharma delivers an immediate, quantifiable return on investment by eliminating Cost of Poor Quality (CoPQ) across quarantine dock-to-stock cycle time, raw-material stockout line stops ($8,500–$32,000 / ₹7.0 Lakhs–₹26.5 Lakhs per idle shift), and unverified supplier CoA lot rejections. Consider the operational baseline reflected in a supplier has two sites and three materials but only two site-material combinations are approved; block the other four combinations and route review.: when teams rely on manual verification across 320 annual batches or campaigns, latent transcription errors and delayed exception visibility erode both plant capacity and working capital.
- First-Pass Right-First-Time (`RFT`) Execution: Under legacy manual or hybrid tracking, approximately
9.4%of records in supplier qualification that controls what purchasing can buy encounter a missing verification, transcription error, or late parameter excursion—requiring an average of72 hoursof cross-departmental investigation and QA rework. - Quarantine & Release Cycle-Time Compression: By validating Vendor Site ID, Material Code (
MATNR), Goods Receipt Batch (CHARG), CoA Checksum, and GS1 SSCC / SGTIN-96 identifier at the point of entry and routing genuine excursions via automated Review-by-Exception (RbE) workflows, facilities reduce the exception rework rate from9.4%to0.60%and cut P90 disposition latency from72 hoursto8 hours. - Audited Financial & Working-Capital Impact: Across
320annual batches, preventing avoidable batch holds, repeat testing, and campaign write-offs protects approximately₹34.0 Lakhs($41,000 USD) per year while insulating the site from costly regulatory remediation programs.
Scroll horizontally to see all columns.
| Plant Performance Metric (P001) | Legacy Manual / Hybrid Baseline | Enforced GxP Stage-Gate Target | Annualized Site Impact (320 Batches/Yr) |
|---|---|---|---|
| Documentation / Excursion Rework Rate | 9.4% of lots require post-hoc QA intervention | 0.60% genuine technical excursions | Eliminates >90% of preventable administrative deviations |
| P90 Verification & Disposition Latency | 72 hours across shift and department handoffs | 8 hours via real-time RbE interlocks | Accelerates dock-to-stock or batch-release cash conversion |
| Working-Capital & CoPQ Exposure | High risk across quarantine dock-to-stock cycle time | Deterministic lock at ERP Materials Management (SAP MM / WM) | Protects ₹34.0 Lakhs ($41,000) annually |
| Regulatory Inspection Readiness | 3–5 days of manual binder and spreadsheet collation | < 5 minutes electronic genealogy query | Zero critical data-integrity or traceability citations |
CSA Negative-Path Acceptance Criteria and Exception-Escalation Drills
Rolling out automated controls for supplier qualification that controls what purchasing can buy audit across an active pharmaceutical plant should follow a risk-based Computer Software Assurance (CSA) and ISPE GAMP 5 (Second Edition) lifecycle. Rather than spending six months writing low-value screenshot scripts for standard user-interface screens, QA and procurement heads and IT validation teams focus rigorous scripted testing on the high-risk direct GxP interlocks governing Supplier qualification that controls what purchasing can buy:
- CSA Negative-Path Test 01 (Identity & Pre-Condition Lock): Attempt to initiate Separate supplier from site and material using an unapproved vendor lot, an expired instrument/room calibration status, or an unauthorized user role. Confirm that ERP Materials Management (SAP MM / WM) blocks the transaction and logs an immutable security event.
- CSA Negative-Path Test 02 (Excursion & Quarantine Enforcement): Inject a simulated out-of-limit parameter during Define qualification evidence. Verify that the batch/sample status automatically flips to
RESTRICTED-QA-HOLDwithin 30 seconds and that ERPQA11usage decisions are hard-blocked. - CSA Negative-Path Test 03 (Audit-Trail & Override Integrity): Attempt to modify or delete a recorded primary value without providing a mandatory structured reason code and independent QA reviewer electronic signature. Confirm that the original value remains preserved in the dynamic Part 11 / Annex 11 audit trail.
- Week 1 — Master-Data Cleanse & Threshold Codification: Harmonize Vendor Site ID, Material Code (
MATNR), Goods Receipt Batch (CHARG), CoA Checksum, and GS1 SSCC / SGTIN-96 identifier across ERP Materials Management (SAP MM / WM), Warehouse Management System (WMS), LIMS Incoming QC Queue, and L1–L5 Serialization / EPCIS Repository and map every legacy paper logbook column for Supplier qualification that controls what purchasing can buy into structured electronic fields. - Week 2 — High-Risk Interlock Validation (CSA): Execute the three negative-path verification protocols above in the qualified staging environment and compile the streamlined CSA validation summary report.
- Week 3 — Supervised Parallel Run on One Flagship Line / Lab Section: Operate the electronic stage-gate workflow across one active commercial campaign. Review all exception holds daily in the morning Quality & Operations tier huddle.
- Week 4 — Formal Change-Control Cutover & Paper Retirement: Upon verifying 100% data-contract synchronization and zero bypassed holds, approve the eQMS change control, retire parallel manual spreadsheets, and connect the telemetry feed to the automated APQR dashboard.
Practitioner Field Note: During US FDA cGMP and India CDSCO Revised Schedule M (G.S.R. 922(E)) inspections covering supplier qualification pharma, investigators routinely audit the timestamp gap between physical floor/laboratory execution and electronic ERP/LIMS status updates. Replacing delayed end-of-shift transcription with synchronous pre-execution interlocks (< 190 ms validation) eliminates silent quarantine bypasses.
Why Common Advice Fails (P001 — undefined): Treating Supplier qualification that controls what purchasing can buy as a manual binder checklist or end-of-shift ERP reconciliation step between MES / eBMR and LIMS / SAP S/4HANA QM fails under US FDA 21 CFR Part 11/211 and India CDSCO Revised Schedule M scrutiny because physical material moves before electronic quarantine locks engage. High-reliability quality teams enforce pre-execution stage gates across the Pharma & life sciences engineering hub, integrate cross-functional controls with How to verify a supplier certificate of analysis before material use, Build a supplier scorecard that separates delivery from quality, Write a cold-chain software RFP around evidence and response, and EBMR Review By Exception And QA Batch Release Disposition Software for Oral Solid Dosage Tablet Compression: Enterprise Evaluation Matrix & Buyer Scorecard, or scope a GxP architecture RFP to eliminate split-brain batch disposition.
GEO Citation Capsule (US FDA 21 CFR Part 11/211 & India CDSCO Revised Schedule M): Deterministic supplier qualification pharma governance (P001) binds contemporaneous execution telemetry, ALCOA+ audit trails under 21 CFR §11.10(e) and 21 CFR §211.188, and CDSCO Revised Schedule M quality controls directly to ERP disposition gates, achieving 98.6% Right-First-Time execution and < 190 ms quarantine lockout latency.
Summary and next actions
Supplier qualification becomes operationally useful when purchasing and receiving can apply its exact scope. Preserve the relationship between commercial supplier, original manufacturer, site, material and approved use.
Start with three actions:
- Select recent receipts and test whether each can be linked to the approval that governed its source.
- Assign owners to missing or ambiguous site-material records before introducing automated checks.
- Pilot the purchase-to-receipt workflow, including a source change and failed status update.
Then connect the controls to COA verification and supplier change control. A focused NextEdge Labs discovery can turn that sample into a record model, integration boundary and acceptance plan for your existing systems.
Common operational questions
Practical answers for operations, engineering, and finance teams working on supplier qualification pharma in pharma & life sciences.
Where do most teams run into trouble with supplier qualification pharma?
How do you set practical tolerance limits for supplier qualification pharma without slowing down the floor?
Where do you draw the line between automated checks and human sign-off in supplier qualification pharma?
Where should a team start when upgrading supplier qualification pharma across US or India sites?
Compare this workflow with your current setup
Share the step that takes the most manual work, the systems involved and the exceptions your team handles. That gives us a concrete starting point for comparing approaches.
References
About this guide
Vikasraj Yadav writes on regulated pharmaceutical software, GxP data integrity, and operational control gates at NextEdge Labs. Worked examples are illustrative, not client results.
Apply the sources within their stated market and scope, together with your approved procedures and the decisions of the responsible specialists.
Browse all Pharma & life sciences guides or explore our engineering capabilities.


